Design verification
Plans, methods, protocols and reports establish whether specified design outputs meet documented design inputs.
Medical devices
Medical device teams must show that design outputs meet design inputs, the finished device satisfies user needs and intended uses, risk controls are verified, and production processes consistently achieve planned results. The strongest document chain preserves those relationships from planning through approval and change.
Industry application concept — device class, market and quality-system scope must be established with the customer.
Where documentation concentrates
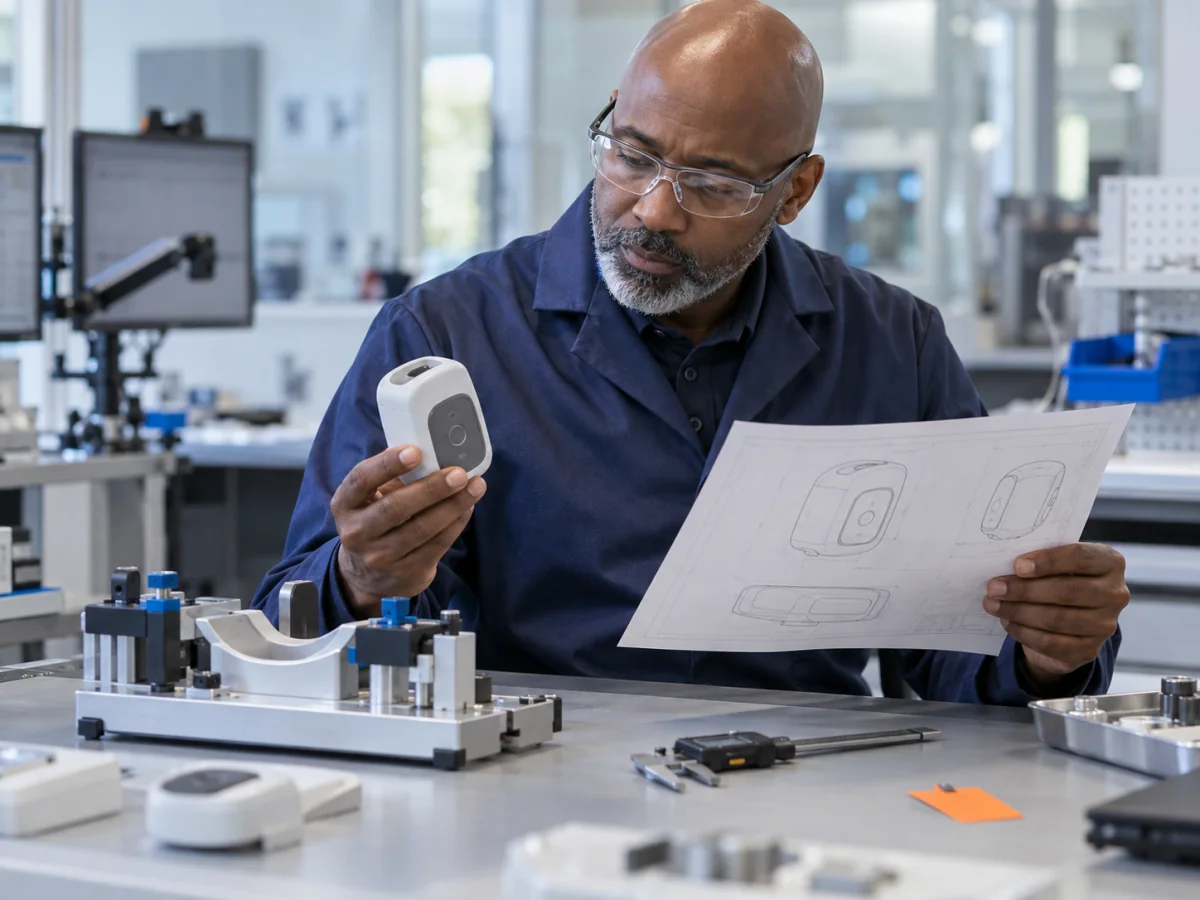
Device documentation is most useful when plans, protocols, results and reports remain traceable to intended use, design inputs, critical characteristics and risk controls.
Plans, methods, protocols and reports establish whether specified design outputs meet documented design inputs.
Evidence addresses user needs and intended uses using production units or justified equivalents under defined conditions.
Hazards, hazardous situations, risk controls, verification evidence and residual-risk conclusions remain connected.
Protocols and reports support processes whose output cannot be fully verified by later monitoring or measurement.
Intended use and risk guide assurance activities for software used in production or the quality management system.
Study plans and reports connect intended users, use environments, critical tasks, endpoints, observations and conclusions.
Representative lifecycle
Verification and validation are not interchangeable. The workflow should preserve the purpose and acceptance logic of each activity.
Users, environments, indications, claims and applicable requirements.
Design inputs, critical characteristics, hazards and risk controls.
Verification, validation, sampling, methods and acceptance criteria.
Independent and qualified reviewers confirm readiness and traceability.
Record configurations, results, anomalies, deviations and source evidence.
Resolve discrepancies and document conclusions against requirements.
Link evidence to the design history, change control and postmarket learning.
Source-to-output map
The platform should help organize evidence without deciding whether a device is safe, effective, validated or releasable.
Engineering, clinical, human-factors, software, quality and regulatory professionals define requirements, approve methods, evaluate anomalies, interpret evidence and authorize release. SmartX supports the document and traceability work surrounding those decisions.
Representative documents
The exact record structure belongs to the manufacturer's quality management system.
Accountable roles
Review routes should reflect the type of device, evidence and risk—not a generic approval list.
Critical reconciliation points
A test result cannot close a requirement or risk control unless the tested article, software build, method and acceptance logic are the ones the approved plan intended.
User needs, design inputs, hazards, risk controls and verification methods should form a visible chain, including the rationale for any requirement that is not directly tested.
Device revision, software build, accessories, fixtures, calibration status, environment, sample rationale and protocol deviation history must identify exactly what evidence was generated.
Failed tests, observations, unresolved anomalies, retesting, concessions and design changes should connect to documented disposition and updates to risk and traceability records.
Define the device family, classification, intended use and target markets; provide approved quality-system templates, one completed evidence chain, risk and traceability formats, source-system locations, naming and configuration rules, and the reviewer and release-authority matrix.
Primary reference points
General educational information only. Device classification, intended use, market and organizational procedures determine applicability. References reviewed October 2026.
Evaluate one evidence chain
A scoped pilot can test template fit, traceability and reviewer controls without changing the manufacturer's approval authority.