Process characterization
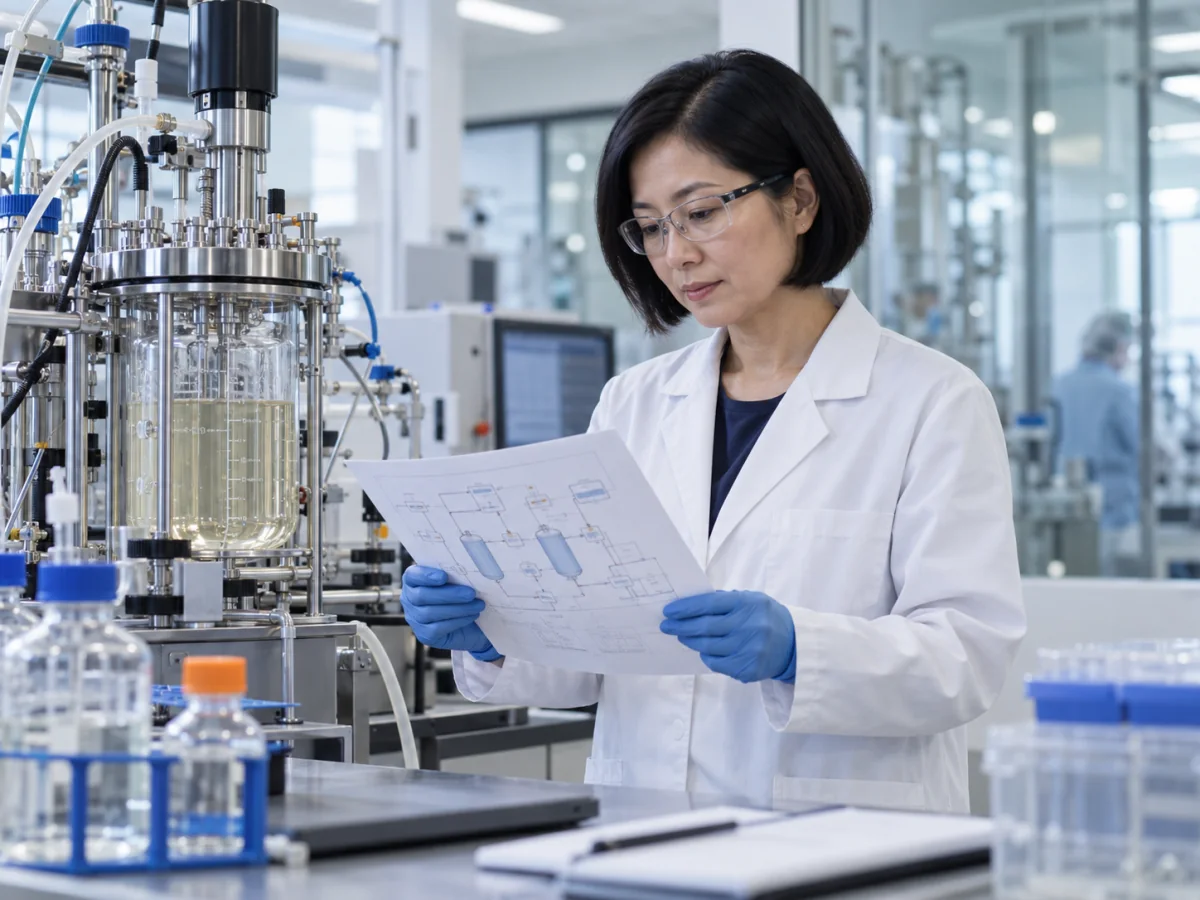
Study plans and reports connect unit operations, parameters, ranges, material attributes and product quality outcomes.
Biotechnology and biologics
Biologics teams coordinate process development, analytical control, manufacturing science, quality and regulatory knowledge across a product whose critical attributes can be sensitive to materials, process conditions and change. The document chain must make the scientific rationale, evidence and decisions easy to follow.
Industry application concept — configuration and validation would be defined with the customer.
Where documentation concentrates
The exact package depends on modality, development stage, jurisdiction and the organization's quality system. These are representative document families.
Study plans and reports connect unit operations, parameters, ranges, material attributes and product quality outcomes.
Procedure development, qualification, validation, transfer and lifecycle records demonstrate fitness for intended use.
Protocols and reports define readiness, acceptance criteria, sampling, execution evidence and continued verification.
Change descriptions, risk assessments, analytical evidence and conclusions support evaluation before and after change.
Protocols, test schedules, data reviews and reports connect quality attributes to storage, shelf life and release controls.
Knowledge-transfer plans, site readiness, method transfer, gaps, responsibilities and closeout evidence support reproducibility.
Representative lifecycle
A comparability, validation or characterization package can use the same governed pattern even when the science differs.
Intended purpose, product stage, change or risk being evaluated.
Product attributes, process data, prior studies, methods and specifications.
Studies, sampling, methods, statistics, criteria and decision rules.
Technical, quality and regulatory roles resolve gaps before work begins.
Results, deviations, metadata and source records remain connected.
Qualified experts assess results against the approved strategy.
Conclusions feed control strategy, change management and lifecycle review.
Source-to-output map
A useful automation boundary begins with approved evidence and ends with a draft that exposes gaps rather than inventing answers.
Process scientists, analytical scientists, biostatisticians, quality and regulatory professionals determine study adequacy, interpret results and approve conclusions. SmartX supports the controlled document workflow around those decisions.
Representative documents
Each customer decides which document types, data sources and approval rules belong in scope.
Accountable roles
Responsibility matrices vary, but the document should make authorship, review, decision ownership and approval explicit.
Critical reconciliation points
Biologics evidence can be internally correct yet still support the wrong conclusion if the product, process, analytical method or change context is misaligned.
Cell substrate or source material, raw-material controls, scale, equipment train, site, process parameters and manufacturing stage must identify the same evaluated process.
Procedure versions, reference standards, system-suitability rules, data-treatment conventions and effective specifications must match the samples and decisions they support.
The change description, affected quality attributes, pre- and post-change evidence, known method limitations, residual uncertainty and follow-up commitments should remain visible together.
Define the modality and lifecycle stage; provide the approved protocol and report templates, one completed example, the source-system inventory, controlled terminology and identifiers, the reviewer and approval matrix, and the rules for handling deviations, changes and unresolved evidence.
Primary reference points
General educational information only. Requirements and accepted approaches depend on product, lifecycle stage, jurisdiction and the organization's quality system. References reviewed October 2026.
Evaluate one document chain
A scoped pilot can establish terminology, traceability, review boundaries and template fit before broader configuration.